Curcumin (Diferuloylmethane) Inhibits DNMTs and HDACs to Restore p53 Tumor Suppressor Signaling
Copy code to add this image to your page
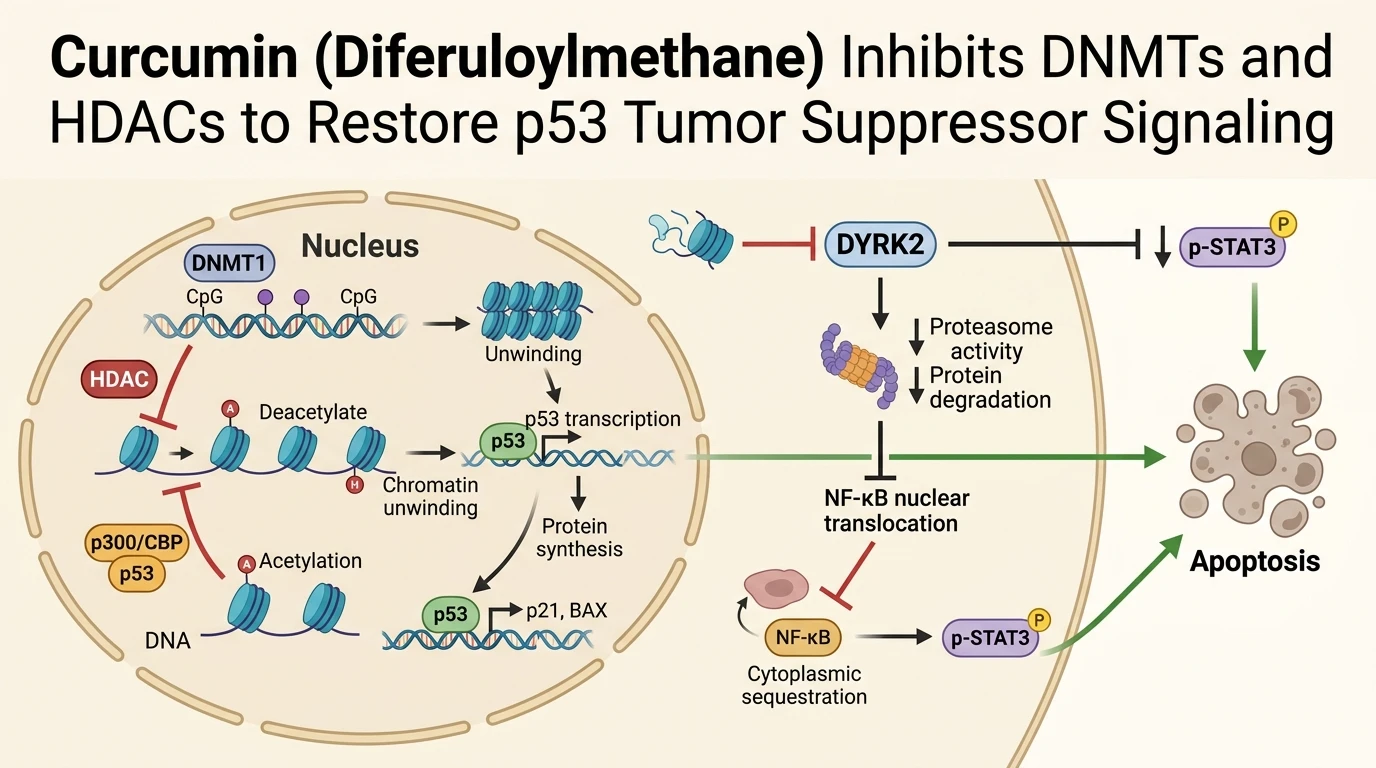
Direct catalytic targeting of epigenetic enzymes offers a therapeutic avenue to reverse aberrant promoter hypermethylation and histone modifications in transformed cellular phenotypes. Curcumin (Diferuloylmethane) acts as a non-nucleoside multi-target epigenetic modifier, direct-binding inhibitor, and transcriptional regulator across diverse neoplastic cell models. Kinetic studies demonstrate that Curcumin (Diferuloylmethane) directly occupies the catalytic pocket of DNA methyltransferase 1 (DNMT1), covalent or non-covalent interactions yielding competitive inhibition against S-adenosyl-L-methionine (SAM). Under tested in vitro assay conditions, this covalent interaction reduces global DNA methylation levels without inducing immediate cytotoxicity. Lowered methyltransferase activity decreases promoter hypermethylation at CpG islands within the TP53 and CDKN2A genomic loci, facilitating transcriptional derepression of p53 and p16INK4a proteins.
Histone modifications represent an interconnected secondary axis of transcriptional control. The small molecule directly binds the C-terminal enzymatic domain of acetyltransferase p300/CBP, covalently modifying cysteine residues within its active site. This reaction prevents acetyl-CoA binding and suppresses histone H3 lysine 9 (H3K9) and H3K14 acetylation profiles. Concurrently, expression levels of class I and class II histone deacetylases (HDACs, including HDAC1, HDAC3, and HDAC8) decrease following treatment at low micromolar concentrations (1–10 µM) in patient-derived cell models. The net alterations in histone acetylation restore a permissive chromatin configuration at tumor suppressor gene promoters while promoting hyperacetylation-induced degradation of oncogenic factors. Investigators at the chromatin biology research center have documented similar multi-layered epigenetic regulatory cascades in mammalian transcriptional control.
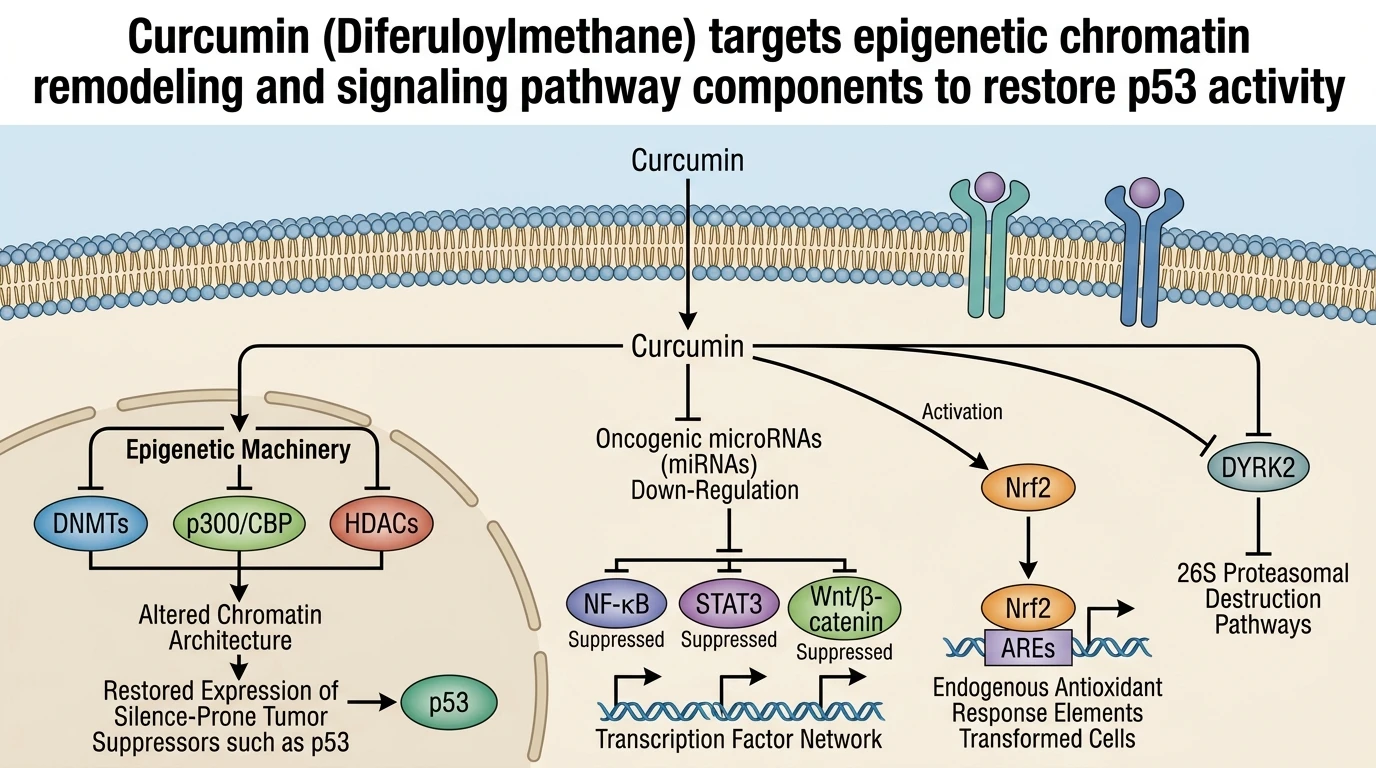
Non-coding RNA profiling reveals additional regulatory nodes sensitive to this polyphenol. The compound selectively upregulates tumor suppressor microRNAs—notably miR-15a, miR-16, and miR-34a—while repressing oncogenic miR-21 and miR-17-92 clusters in human breast and colon carcinoma cell lines. Amplified miR-34a expression reinforces p53 signaling loops, promoting down-regulation of anti-apoptotic BCL2 and cyclin D1 translation. In parallel, decreased miR-21 expression limits phosphatase and tensin homolog (PTEN) silencing, restoring intracellular PIP3/AKT signaling control.
Beyond classical epigenetic regulatory enzymes, direct structural interactions alter protein turnover and stress response cascades. Kinase profiling demonstrates competitive binding to dual-specificity tyrosine-regulated kinase 2 (DYRK2). Selective inhibition of DYRK2 prevents catalytic phosphorylation of the 26S proteasome subunit RPT6 at Ser120. Consequently, proteasomal assembly is impaired, suppressing protein degradation pathways essential for rapid tumor growth. Parallel induction of nuclear factor erythroid 2-related factor 2 (Nrf2) nuclear translocation occurs via Keap1 modification. Nrf2 binds antioxidant response elements (ARE), upregulating heme oxygenase-1 (HO-1) and NAD(P)H:quinone oxidoreductase 1 (NQO1) expression.
Downstream signal transduction networks undergo marked inhibition. Attenuation of IκB kinase (IKK) complex phosphorylation blocks IκBα degradation, trapping nuclear factor kappa B (NF-κB) p65/p50 heterodimers in the cytoplasm. Simultaneously, Signal Transducer and Activator of Transcription 3 (STAT3) phosphorylation at Tyr705 declines within hours of exposure in preclinical models. Wnt/β-catenin signaling decreases through enhanced glycogen synthase kinase 3 beta (GSK-3β) activation and downstream β-catenin degradation. These concurrent pathway disruptions culminate in G2/M phase cell cycle arrest, caspase-3 activation, and execution of intrinsic apoptosis in transformed cells.
Pharmacological Epigenetics and Multi-Target Signaling Modulation
Copy code to add this image to your page
Curcumin (Diferuloylmethane) functions as a broad epigenetic and transcriptional regulator by inhibiting DNMT1, p300/CBP HAT, and class I HDACs. These enzymatic perturbations decrease promoter hypermethylation while enhancing permissive histone acetylation, directly reactivating TP53 and suppressing miR-21. Simultaneous inhibition of DYRK2 limits 26S proteasome activity, stabilizing IκBα and blocking NF-κB p65 nuclear import. Concurrent downregulation of STAT3 and Wnt/β-catenin pathways combines with Nrf2-mediated antioxidant activation to enforce cell cycle arrest and intrinsic apoptosis in transformed cell models.
DNMT1 Catalytic Pocket Inhibition Restores p53 Expression
Competitive binding within the DNMT1 catalytic center inhibits covalent methyl transfer to CpG dinucleotides. Under tested in vitro assay conditions, treatment with 5 µM Curcumin (Diferuloylmethane) reduces locus-specific promoter methylation by nearly forty percent. Localized demethylation reactivates silent TP53 promoters within hours. Transcriptional reactivation followed immediately after. Reactivated p53 drives transcription of p21CIP1/WAF1, arresting cells at the G1/S transition checkpoint in human colorectal carcinoma models. Downstream apoptotic signaling pathways activated simultaneously. Restored p53 also induces PUMA expression, triggering mitochondrial outer membrane permeabilization and cytochrome c release.
DYRK2 Proteasomal Suppression Blocks Oncogenic NF-kB Activation
Direct binding to DYRK2 disrupts 26S proteasome assembly by blocking RPT6 phosphorylation at Ser120. Proteasomal degradation rates dropped significantly. In human triple-negative breast cancer models, reduced proteasomal activity prevents the rapid turnover of IκBα inhibitor proteins. Cytoplasmic retention of NF-κB p65 ensues. Nuclear translocation stopped abruptly thereafter. Without nuclear p65 dimers, transcription of anti-apoptotic targets including BCL-XL, XIAP, and COX-2 drops below detection limits in vitro. Unphosphorylated STAT3 levels remained stable while active p-STAT3 Tyr705 content fell sharply. Curcumin (Diferuloylmethane) administration thus decouples inflammatory transcriptional responses from survival signaling nodes in malignant cells.
Histone Deacetylase Suppression Promotes Hyperacetylation-Induced Apoptosis
Inhibition of class I HDACs elevates histone H3 and H4 acetylation state across promoter regions. Chromatin accessibility expanded within two hours. Hyperacetylated histone tails alter nucleosome density, exposing promoter elements to transcription factor complexes. Cell cycle progression ceased rapidly. Increased H3K9 acetylation correlates with attenuated β-catenin expression in SW480 colon adenocarcinoma cells. Nuclear accumulation of β-catenin declined markedly. Lowered Wnt pathway activity suppresses MYC and CCND1 transcription, enforcing G2/M arrest and activating executioner caspase-3 cascades under experimental conditions.